

Automated Quantum Algorithm Discovery for Quantum Chemistry
Quantinuum, the world's leading quantum computing company, and Hiverge, a Cambridge-based startup developing an engine for automated algorithm discovery, have initiated a collaboration on quantum algorithm discovery.
Authors:
Quantinuum (alphabetical order): Eric Brunner, Steve Clark, Fabian Finger, Gabriel Greene-Diniz, Pranav Kalidindi, Alexander Koziell-Pipe, David Zsolt Manrique, Konstantinos Meichanetzidis, Frederic Rapp
Hiverge (alphabetical order): Alhussein Fawzi, Hamza Fawzi, Kerry He, Bernardino Romera Paredes, Kante Yin
What if every quantum computing researcher had an army of students to help them write efficient quantum algorithms? Large Language Models are starting to serve as such a resource.
Quantinuum’s processors offer world-leading fidelity, and recent experiments show that they have surpassed the limits of classical simulation for certain computational tasks, such as simulating materials. However, access to quantum processors is limited and can be costly. It is therefore of paramount importance to optimise quantum resources and write efficient quantum software. Designing efficient algorithms is a challenging task, especially for quantum algorithms: dealing with superpositions, entanglement, and interference can be counterintuitive.
To this end, our joint team used Hiverge’s AI platform for automated algorithm discovery, the Hive, to probe the limits of what can be done in quantum chemistry. The Hive generates optimised algorithms tailored to a given problem, expressed in a familiar programming language, like Python. Thus, the Hive’s outputs allow for increased interpretability, enabling domain experts to potentially learn novel techniques from the AI-discovered solutions. Such AI-assisted workflows lower the barrier of entry for non-domain experts, as an initial sketch of an algorithmic idea suffices to achieve state-of-the-art solutions.
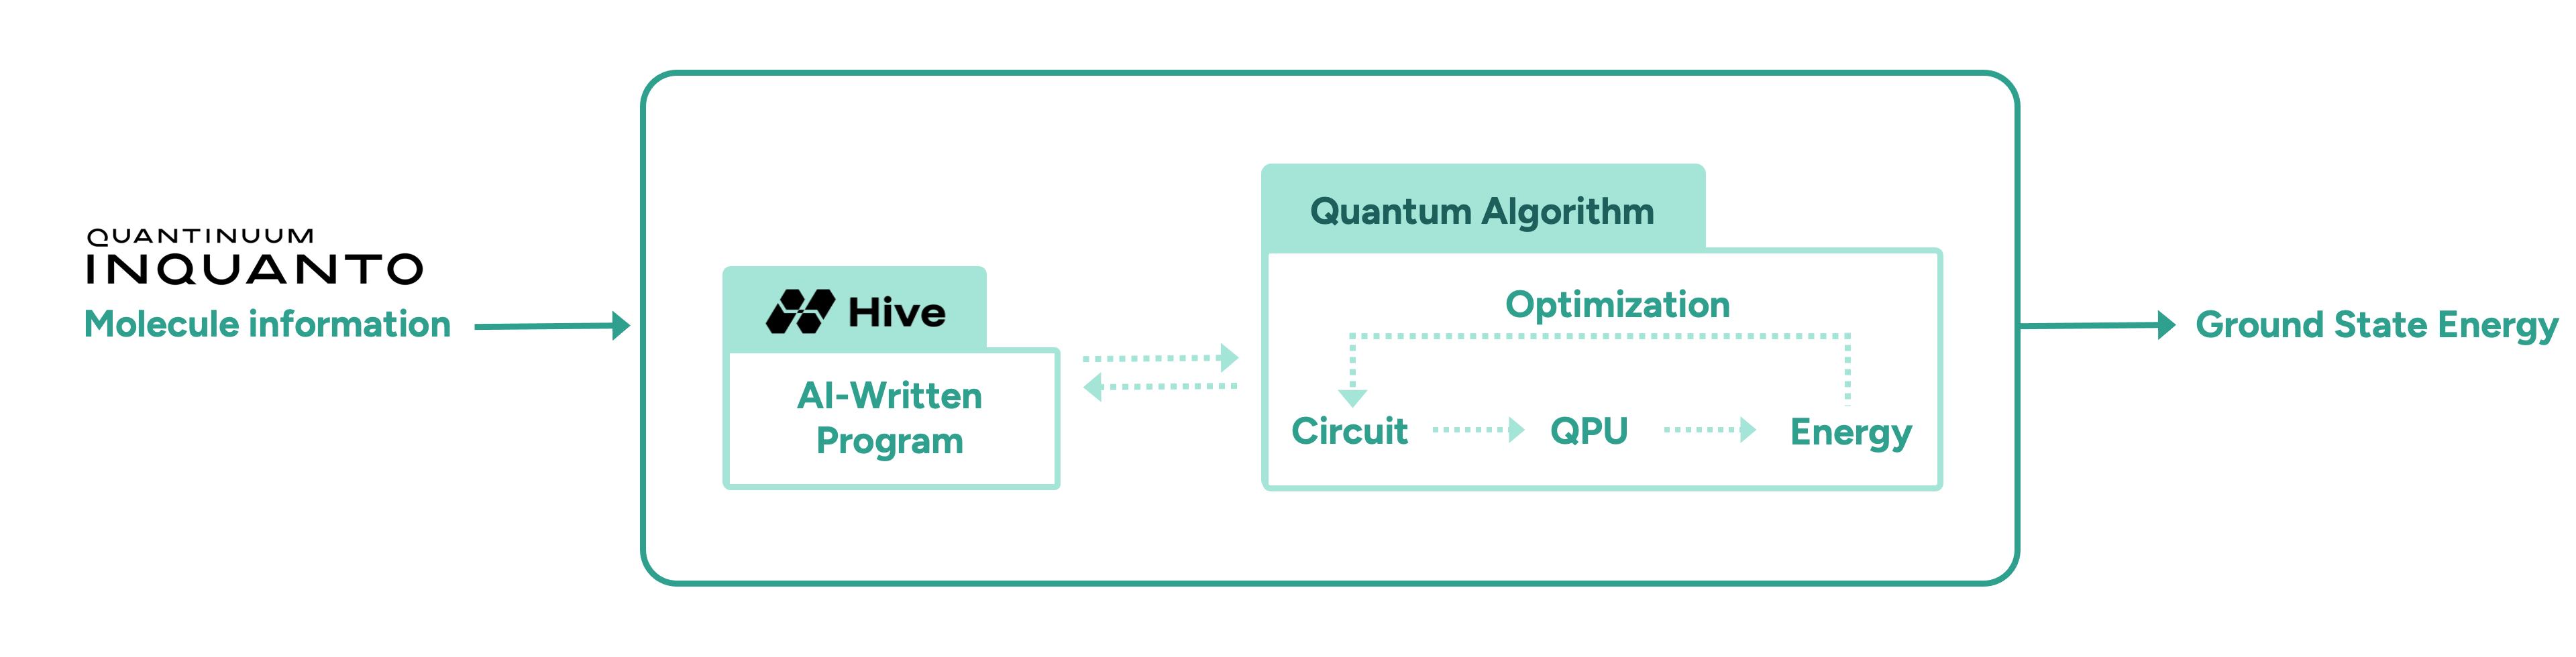
In this initial proof-of-concept study, we demonstrate the advantage of AI-driven algorithmic discovery of efficient quantum heuristics in the context of quantum chemistry, in particular the electronic structure problem. Our early explorations show that the Hive can start from a naïve and simple problem statement and evolve a highly optimised quantum algorithm that solves the problem, reaching chemical precision for a collection of molecules. Our high-level workflow is shown in Figure 1. Specifically, the quantum algorithm generated by the Hive achieves a reduction in the quantum resources required by orders of magnitude compared to current state-of-the-art quantum algorithms. This promising result may enable the implementation of quantum algorithms on near-term hardware that was previously thought impossible due to current resource constraints.
The Electronic Structure Problem in Quantum Chemistry
The electronic structure problem is central to quantum chemistry. The goal is to prepare the ground state (the lowest energy state) of a molecule and compute the corresponding energy of that state to chemical precision or beyond. Classically, this is an exponentially hard problem. In particular, classical treatments tend to fall short when there are strong quantum effects in the molecule, and this is where quantum computers may be advantageous.
The paradigm of variational quantum algorithms is motivated by near-term quantum hardware. One starts with a relatively easy-to-prepare initial state. Then, the main part of the algorithm consists of a sequence of parameterised operators representing chemically meaningful actions, such as manipulating electron occupations in the molecular orbitals. These are implemented in terms of parameterised quantum gates. Finally, the energy of the state is measured via the molecule’s energy operator, the “Hamiltonian”, by executing the circuit on a quantum computer and measuring all the qubits on which the circuit is implemented. Taking many measurements, or “shots”, the energy is estimated to the desired precision. The ground state energy is found by iteratively optimising the parameters of the quantum circuit until the energy converges to a minimum value. The general form of such a variational quantum algorithm is illustrated in Figure 2.
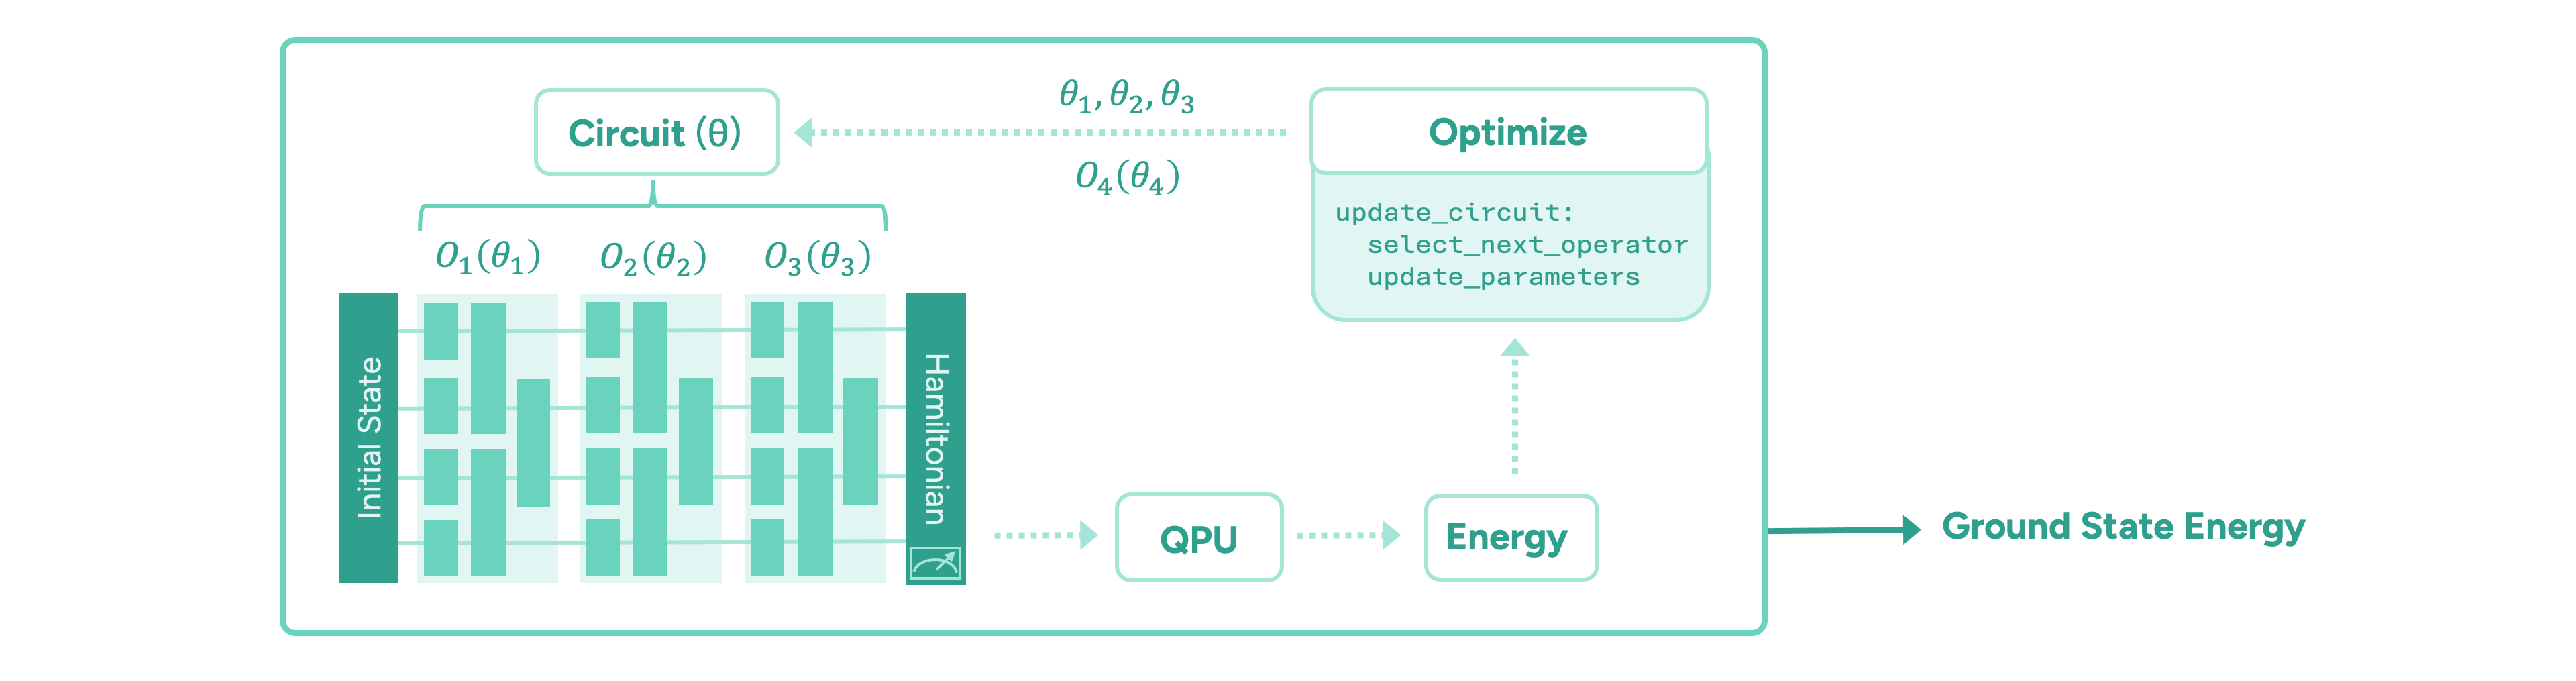
The main challenge in these frameworks is to design an appropriate quantum circuit architecture, i.e. find an efficient sequence of operators, and an efficient optimisation strategy for its parameters. It is important to minimise the number of quantum operations in any given circuit, as each operation is inherently noisy and the algorithm’s output degrades exponentially. Another important quantum resource to be minimised is the total number of circuits that need to be evaluated to compute the energy values during the optimisation of the circuit parameters, which is time-consuming.
To meet these challenges, we task the Hive with designing a variational quantum algorithm to solve the ground state problem, following the workflow shown in Figure 1. The Hive is a distributed evolutionary process that evolves programs. It uses Large Language Models to generate mutations in the form of edits to an entire codebase. This genetic process selects the fittest programs according to how well they solve a given problem. In our case, the role of the quantum computer is to compute the fitness, i.e., the ground state energy. Importantly, the Hive operates at the level of a programming language; it readily imports and uses all known libraries that a human researcher would use, including Quantinuum’s quantum chemistry platform, InQuanto. In addition, the Hive can accept instructions and requests in natural language, increasing its flexibility. For example, we encouraged it to seek parameter optimisation strategies that avoid estimating gradients, as this incurs significant overhead in terms of circuit evaluations. Intuitively, the interaction between a human scientist and the Hive is analogous to a supervisor and a group of eager and capable students: the supervisor provides guidance at a high level, and the students collaborate and flesh out the general idea to produce a working solution that the supervisor can then inspect.
We find that from an extremely basic starting point, consisting of a skeleton for a variational quantum algorithm, the Hive can autonomously assemble a bespoke variational quantum algorithm, which we call Hive-ADAPT. Specifically, the Hive evolves heuristic functions that construct a circuit as a sequence of quantum operators and optimise its parameters. Remarkably, the Hive converged on a structure resembling the current state-of-the-art, ADAPT-VQE. Crucially, however, Hive-ADAPT substantially outperforms this baseline, delivering significant improvements in chemical precision while reducing quantum resource requirements.
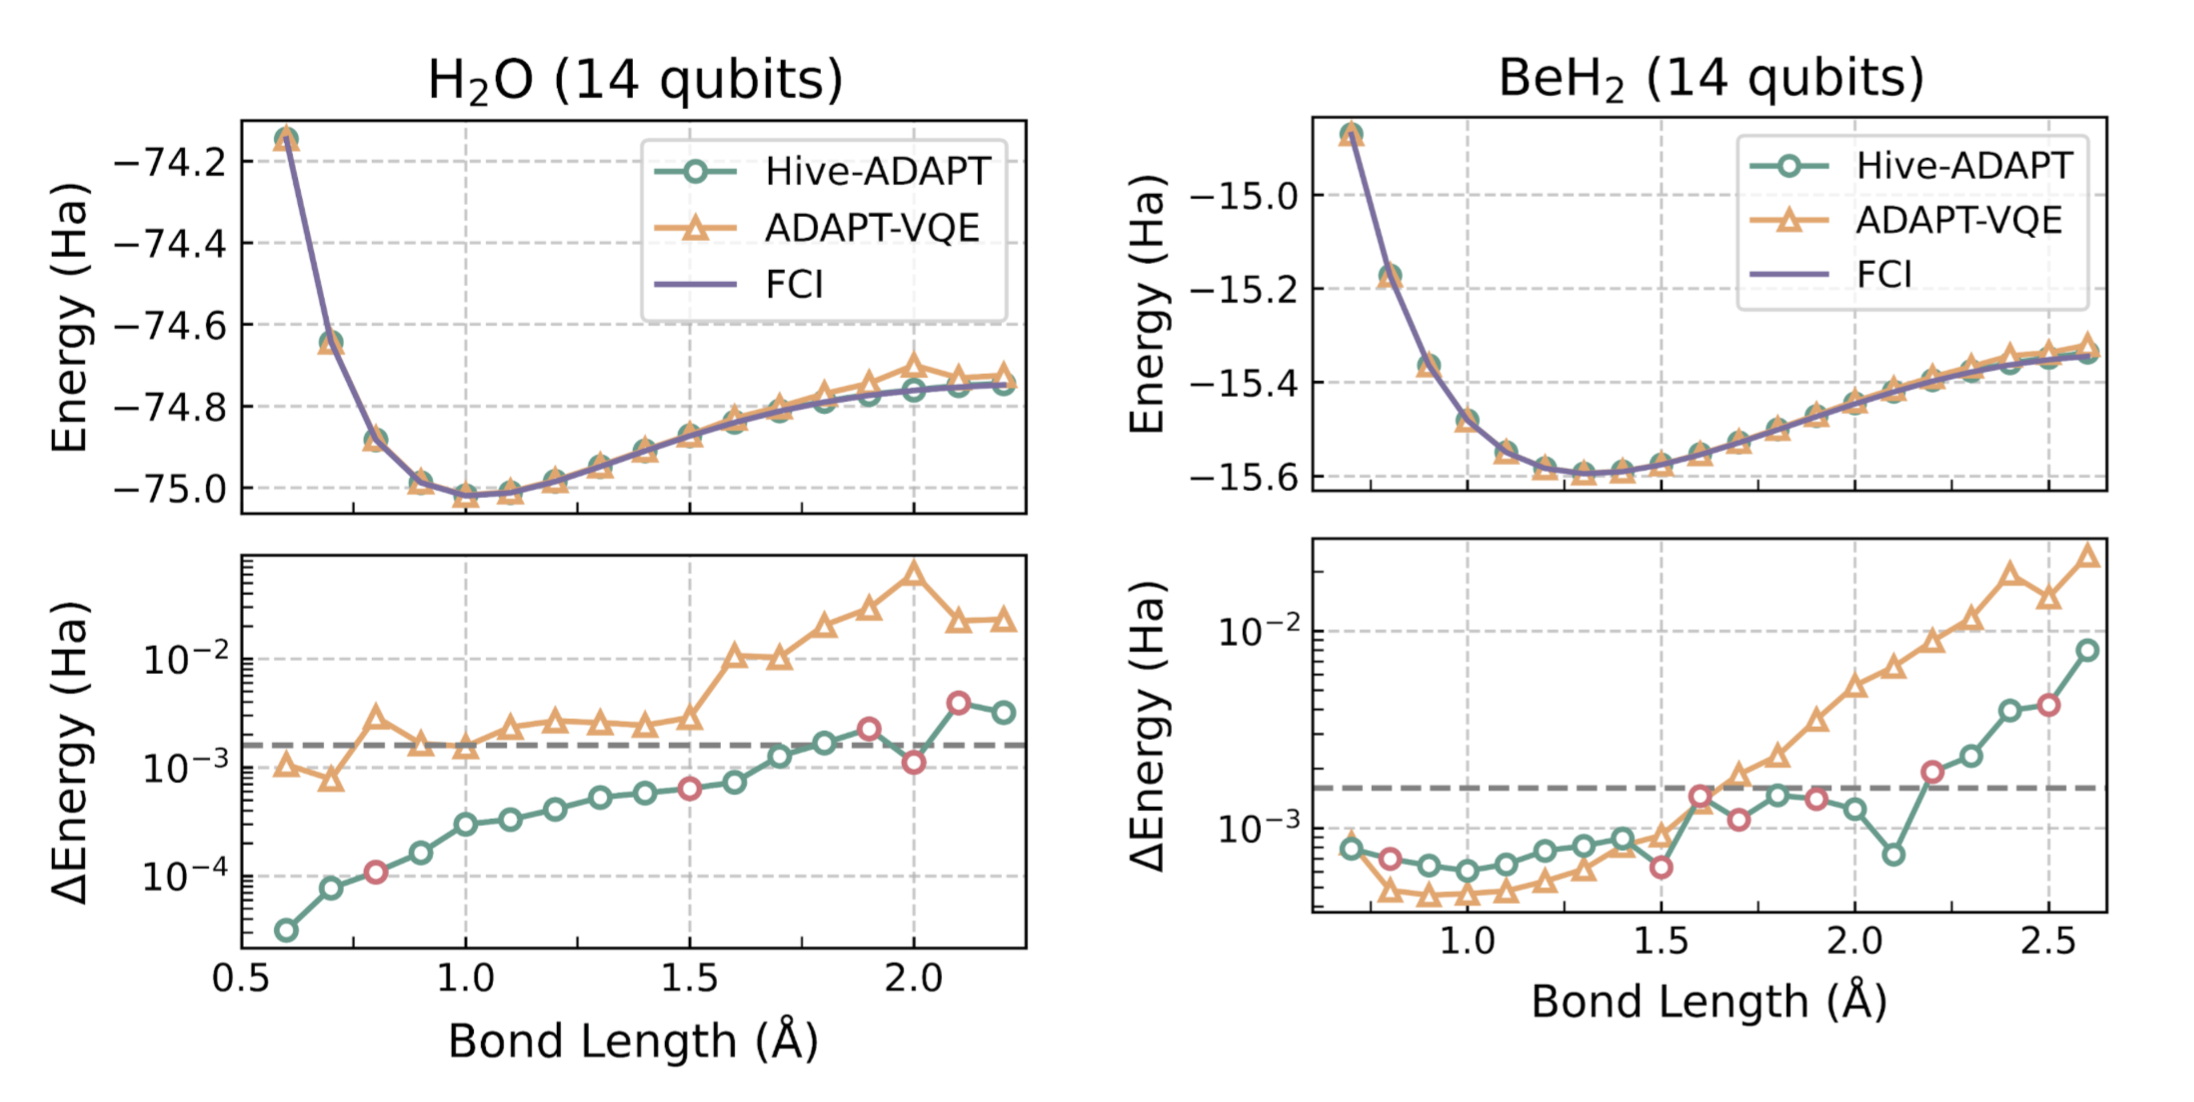
A molecule’s ground state energy varies with the distances between its atoms, called the “bond length”. For example, for the molecule H2O, the bond length refers to the length of the O-H bond. The Hive was tasked with developing an algorithm for a small set of bond lengths and reaching chemical precision, defined as within 1.6e-3 Hartree (Ha) of the ground state energy computed with the exact Full Configuration Interaction (FCI) algorithm. As we show in Figure 3, remarkably, Hive-ADAPT achieves chemical precision for more bond lengths than ADAPT-VQE. Furthermore, Hive-ADAPT also reaches chemical precision for other “unseen” bond lengths, showcasing the generalisation ability of the evolved quantum algorithm. Our results were obtained from classical simulations of the quantum algorithms, where we used NVIDIA CUDA-Q to leverage the parallelism enabled by GPUs. Further, relative to ADAPT-VQE, Hive-ADAPT exhibits one to two orders of magnitude reduction in quantum resources, such as the number of circuit evaluations and the number of operators used to construct circuits, which is crucial for practical implementations on actual near-term processors.
For molecules such as BeH2 at large Be-H bond lengths, a complex initial state is required for the algorithm to be able to reach the ground state using the available operators. Even in these cases, by leveraging an efficient state preparation scheme implemented in InQuanto, the Hive evolved a dedicated strategy for the preparation of such a complex initial state, given a set of basic operators to achieve the desired chemical precision.
To validate Hive-ADAPT under realistic conditions, we employed Quantinuum’s H2 Emulator, which provides a faithful classical simulator of the H2 quantum computer, characterised by a 1.05e-3 two-qubit gate error rate. Leveraging the Hive's inherent flexibility, we adapted the optimisation strategy to explicitly penalise the number of two-qubit gates—the dominant noise source on near-term hardware—by redefining the fitness function. This constraint guided the Hive to discover a noise-aware algorithm capable of constructing hardware-efficient circuits. We subsequently executed the specific circuit generated by this algorithm for the LiH molecule at a bond length of 1.5 Å with the Partition Measurement Symmetry Verification (PMSV) error mitigation procedure. The resulting energy of -7.8767 ± 0.0031 Ha, obtained using 10,000 shots per circuit with a discard rate below 10% in the PMSV error mitigation procedure, is close to the target FCI energy of -7.8824 Ha and demonstrates the Hive's ability to successfully tailor algorithms that balance theoretical accuracy with the rigorous constraints of hardware noise and approach chemical precision as much as possible with current quantum technology.
For illustration purposes, we show an example of an elaborate code snippet evolved by the Hive starting from a trivial version:
Quantinuum’s in-house quantum chemistry expert, Dr. David Zsolt Manrique, commented,
“I found it amazing that the Hive converged to a domain-expert level idea. By inspecting the code, we see it has identified the well-known perturbative method, ‘MP2’, as a useful guide; not only for setting the initial circuit parameters, but also for ordering excitations efficiently. Further, it systematically and laboriously fine-tuned those MP2-inspired heuristics over many iterations in a way that would be difficult for a human expert to do by hand. It demonstrated an impressive combination of domain expertise and automated machinery that would be useful in exploring novel quantum chemistry methods.”
Looking to the Future
In this initial proof-of-concept collaborative study between Quantinuum and Hiverge, we demonstrate that AI-driven algorithm discovery can generate efficient quantum heuristics. Specifically, we found a great reduction in quantum resources, which is impactful for quantum algorithmic primitives that are frequently reused. Importantly, this approach is highly flexible; it can accommodate the optimisation of any desired quantum resource, from circuit evaluations to the number of operations in a given circuit. This work opens a path toward fully automated pipelines capable of developing problem-specific quantum algorithms optimised for NISQ as well as future hardware.
An important question for further investigation regards transferability and generalisation of a discovered quantum solution to other molecules, going beyond the generalisation over bond lengths of the same molecule that we have already observed. Evidently, this approach can be applied to improving any other near-term quantum algorithm for a range of applications from optimisation to quantum simulation.
We have already demonstrated an error-corrected implementation of quantum phase estimation on quantum hardware, and an AI-driven approach promises further hardware-tailored improvements and optimal use of quantum resources. Beyond NISQ, we envision that AI-assisted algorithm discovery will be a fruitful endeavour in the fault-tolerant regime, as well, where high-level quantum algorithmic primitives (quantum fourier transform, amplitude amplification, quantum signal processing, etc.) are to be combined optimally to achieve computational advantage for certain problems.
Notably, we’ve entered an era where quantum algorithms can be written in high-level programming languages, like Quantinuum’s Guppy, and approaches that integrate Large Language Models directly benefit. Automated algorithm discovery is promising for improving routines relevant to the full quantum stack, for example, in low-level quantum control or in quantum error correction.
About Quantinuum
Quantinuum, the world’s largest integrated quantum company, pioneers powerful quantum computers and advanced software solutions. Quantinuum’s technology drives breakthroughs in materials discovery, cybersecurity, and next-gen quantum AI. With over 500 employees, including 370+ scientists and engineers, Quantinuum leads the quantum computing revolution across continents.
- Our team, in partnership with HQS Quantum Simulations, just published what we believe to be the most accurate1 large-scale quantum simulation of Nuclear Magnetic Resonance (NMR) to date, pointing the way to new frontiers in computational chemistry.
- NMR is a widely used analytical technique that can require significant computational costs to interpret, costs that can balloon quickly on classical systems.
- In this result, the joint team ran an end-to-end simulation on System Model H2, reproducing key spectral features that previous demonstrations were unable to capture.
Nuclear magnetic resonance (NMR) spectroscopy is one of the most powerful analytical techniques in modern science. From identifying drug candidates to understanding battery materials, it allows researchers to probe the local atomic structure of molecules and materials with remarkable precision.
Interpreting NMR experiments often requires simulations that are just as challenging as the experiments themselves. As the number of interacting nuclear spins grows, the computational cost of simulating these systems increases exponentially on classical computers. Quantum computers, which naturally represent and evolve quantum states, offer a fundamentally different approach.
In our latest work, we demonstrate the most accurate large-scale digital NMR simulation performed on quantum hardware to date. Using Quantinuum's System Model H2, we carried out an end-to-end simulation of a classically challenging NMR experiment, reproducing key spectral features that previous hardware demonstrations were unable to capture.
Why simulate NMR?
NMR is a cornerstone of chemical analysis. Researchers use it to determine molecular structures, characterize new compounds, and study how atoms interact with one another.
These capabilities make NMR indispensable across industry. In pharmaceutical research, NMR helps identify and characterize drug candidates. In materials science, it reveals the local atomic environments that determine material properties. Battery researchers, for example, use NMR to study cathode materials such as lithium cobalt oxide, allowing them to monitor how the material changes during charging and discharging and ultimately improve battery performance.
In many cases, the experimental spectrum is only part of the story. Simulations help scientists interpret complex spectra by revealing which atomic interactions give rise to the observed peaks. They provide the link between an experimental measurement and the underlying molecular structure.
The classical challenge and the quantum solution
The difficulty lies in the physics.
An NMR experiment measures the dynamics of interacting nuclear spins. Every additional spin rapidly increases the size of the quantum state that must be represented (in the case of a spin-½ particle, every spin doubles the state). As a result, exact classical simulations become exponentially more expensive as the system size grows.
Today's best classical methods are remarkably sophisticated. Exact simulations are typically limited to systems of around 25 interacting spins, while advanced approximation techniques can often extend calculations to roughly 40–50 spins for many practical problems.
Those approximations have made classical NMR software extraordinarily effective for conventional liquid-state spectroscopy. In fact, our collaborators on this project are developing one of the leading classical simulation packages and noted that, despite the success of our quantum experiment, the current spectral resolution is still insufficient for routine use by practicing spectroscopists. Resolving the fine structure needed for many real-world analyses would require substantially longer simulations—and therefore much deeper quantum circuits than current hardware can yet support.
This highlights both the progress and the remaining challenge. Quantum computers are beginning to produce meaningful NMR spectra, but practical utility will require larger quantum computers and increased simulation depth.
In the longer term, interacting spin systems are among the most natural applications for quantum computers. Instead of storing the exponentially large quantum state explicitly, a quantum computer represents it directly in its physical qubits. For spin-½ nuclei, the mapping is particularly efficient: each nuclear spin corresponds directly to a single qubit. Higher-spin nuclei require only a small number of additional qubits.
This does not eliminate every computational challenge—longer simulations still require deeper quantum circuits—but it avoids the exponential memory bottleneck that limits classical simulation.
For NMR, this makes quantum simulation an especially compelling long-term application.
Our experiment
Working with collaborators at HQS Quantum Simulations, we implemented a complete quantum workflow for simulating an NMR experiment on Quantinuum's H2 trapped-ion quantum computer.
Rather than stopping at Hamiltonian simulation alone, we reproduced the entire computational pipeline:
- constructing the nuclear spin Hamiltonian,
- compiling efficient quantum circuits,
- simulating the spin dynamics through Trotterized real-time evolution,
- applying targeted error-suppression techniques,
- reconstructing the free induction decay, and
- generating the final NMR spectrum through Fourier analysis.
The benchmark molecule, 1,2-di-tert-butyl-diphosphane, is a well-known challenge for NMR simulation. After applying hardware-efficient model reduction, we simulated an effective 21-spin Hamiltonian using a 42-qubit (21 system qubits and 21 ancilla qubits) computation on System Model H2.
Our deepest circuits reached over 1,400 two-qubit gates and simulated 70 Trotter steps, corresponding to approximately 29 ms of physical evolution time.
Most importantly, the resulting spectrum reproduced the key benchmark features of the classical reference calculation, including a characteristic double-peak structure that previous quantum hardware demonstrations had failed to recover.
This represents the most complete hardware demonstration of digital NMR simulation reported to date.
Why trapped ions mattered
Achieving this result depended not only on the quantum algorithm but also on the underlying hardware.
Large Hamiltonian simulations require long, high-fidelity quantum circuits. Quantinuum's trapped-ion architecture provides all-to-all qubit connectivity, high-fidelity gates, and effective error-suppression techniques that allowed us to preserve the spectroscopic features throughout the computation.
The close agreement between hardware, emulator, and classical reference calculations demonstrates that deep quantum simulations of chemically meaningful systems are becoming increasingly feasible on today's hardware.
Looking ahead
To make quantum NMR genuinely useful for practicing spectroscopists, future systems will need substantially deeper circuits.
Our experiment simulated approximately 29 ms of evolution time, producing spectral features with a resolution of roughly 0.07 ppm, a very promising start.
Reaching that scale will require continued improvements in hardware fidelity, error correction, and quantum algorithms.
Nevertheless, this work demonstrates that digital quantum simulation of realistic NMR experiments is no longer purely theoretical. It establishes a practical end-to-end workflow, validates key algorithmic techniques, and shows that quantum computers can already reproduce chemically meaningful spectral signatures.
As quantum hardware continues to improve, applications such as molecular spectroscopy, materials characterization, and chemical simulation are becoming increasingly realistic targets for practical quantum computing.
1 Based on a study of existing literature
- In a recent independent study by the Julich Supercomputing Center, Quantinuum’s trapped-ion systems outperformed superconducting processors in tests of fault tolerant operations.
- Testing small chunks of circuits that would be used in larger QEC workflows, Quantinuum’s Helios demonstrated an approximately 10x lower effective hardware error compared to superconducting systems.
- Quantinuum’s flexible connectivity enabled tests across three error-correcting code families, compared to just one for superconducting systems, illustrating the expanding options for advancing fault tolerance with our QCCD architecture.
The demands of fault tolerance are bringing Quantinuum’s advantages into sharper focus.
In a new independent study from the Jülich Supercomputing Centre, Quantinuum’s Helios demonstrated an approximately order-of-magnitude advantage over superconducting hardware in tests of operations essential to quantum error correction. More precisely, Helios’ effective hardware error was approximately 13 times lower at 30 data qubits and eight times lower at 50 data qubits than the comparable superconducting result.
The advantage extended to architectural flexibility. Quantinuum’s systems supported tests across three error-correcting code families, while connectivity and control restrictions limited implementation to only one on the superconducting devices evaluated.
These findings build on earlier independent research by the same organization highlighting Quantinuum’s physical-level performance. As we have argued, the NISQ era is coming to an end. The demands of error correction are bringing our architectural differences into sharper focus—and this study provides further evidence of Quantinuum’s advantage.
Error correction exposes performance gaps
A fault-tolerant quantum computer must repeatedly detect and correct errors to protect the computation in progress. That requires reliable mid-circuit measurements, conditional operations, and real-time coordinated scheduling.
In a direct comparison, introducing mid-circuit measurement caused substantially greater degradation on the superconducting processors compared to the Quantinuum systems.
That difference matters. Mid-circuit measurement must be repeated throughout fault-tolerant computations. Their (potential) performance penalty directly affects how much computation a machine can sustain.
Helios demonstrated strong performance under those demands, extending its advantage beyond the physical layer into circuits exercising essential error-correction capabilities.
Connectivity expands the options
The study also highlighted how architectural restrictions affect which error-correction structures hardware can implement directly.
On Quantinuum’s systems, researchers ran tests exploring the surface-code, triangular color-code, and a bivariate-bicycle qLDPC code. In contrast, the superconducting devices evaluated were only able to explore the surface code due to device constraints.
This is because Quantinuum’s QCCD architecture offers greater flexibility: mobile qubits enable codes that require higher connectivity than is allowed on traditional superconducting processors. Ultimately, this gives researchers more freedom to explore multiple codes—and more opportunities to reduce the qubit and time overheads of fault tolerance.
Building on an architectural advantage
The Jülich study provides independent evidence that Quantinuum’s architectural choices deliver advantages for operations essential to fault tolerance. It also reveals performance and implementation gaps in the superconducting systems tested.
We intend to extend that lead. Our investments in fidelity, flexible connectivity, and integrated control are foundations for increasingly capable machines.
As error-correction workloads become more demanding, those capabilities become more consequential. Quantinuum is building to meet that challenge—and to keep raising the performance bar.
Thirty participants gathered at Newnham College, Cambridge, for an intensive five-day programme exploring how quantum computers can be used to model chemical systems.
By: Duncan Gowland, Senior Advanced R&D Scientist at Quantinuum
For 30 researchers who gathered at Newnham College, Cambridge this September, that question shaped an intensive five-day programme.
Quantinuum’s first InQuanto Summer School, supported by the UK’s National Quantum Computing Centre (NQCC), combined lectures, hands-on workshops, and mini-projects to help participants connect fundamental concepts with practical research. By the end of the week, participants were applying those ideas to problems ranging from molecular electronic structure to quantum state-preparation circuits.
Applying quantum computing to chemistry can be challenging because it draws on expertise from a broad range of areas, including electronic structure theory, quantum algorithms, software, hardware, noise modelling, and resource estimation. Researchers often enter the field with deep knowledge of one or two of these areas but less familiarity with the others.
A key objective of the course was to help researchers from computational chemistry and quantum computing build stronger foundations, develop a shared language, and deepen their understanding of adjacent disciplines.
For researchers beginning work at this intersection, that shared understanding can make it easier to identify where to start, which assumptions to challenge, and when to seek expertise from another discipline.
Bringing Theory and Practice Together
The programme began with lectures on the foundations of quantum computing, followed by pen-and-paper and computational exercises in the afternoon. On Day 2, lectures and workshops introduced the electronic structure problem: how quantum chemists describe the behaviour of electrons in molecules.
Students then brought these two foundations together by studying fermion-to-qubit mappings, which translate chemistry problems into a form that quantum computers can process, before moving on to the measurement of chemistry observables and the construction of quantum algorithms.
Learners were given free access to InQuanto, Quantinuum’s quantum chemistry software platform designed to accelerate research in fields such as chemistry and condensed matter physics using quantum computers. InQuanto supported the lectures as a broad, well-documented, and thoroughly tested platform that enabled students to explore and reinforce complex concepts through hands-on experience.
Later in the week, attention shifted to state-of-the-art considerations: how to build useful chemical models, make the best use of current quantum devices, and think about how quantum algorithms for chemistry might develop over the next five to ten years.
The students, who travelled from around the world to Cambridge, brought a remarkable range of backgrounds and experience. The material was pitched at roughly first-year doctoral level, for participants with at least a year of experience in one core area and some Python skills. Even with this experience, participants benefited from exposure to other disciplines and from the practical expertise of Quantinuum’s quantum chemistry team—which brings years of experience working with industrial including BMW Group, TotalEnergies, NVIDIA, and Pfizer.
Throughout the week, participants were highly engaged in lectures and brought a thoughtful, collaborative approach to the workshop exercises. Discussions continued throughout the week, from coffee breaks to lunches and dinners, creating valuable opportunities to exchange ideas and experiences.
A particular highlight was the mini-project work. In just a day and a half, participants tackled a wide range of problems and produced impressive prototype solutions. The projects concluded with a poster-style session, where the quality of discussion was exceptional. Examples included state-preparation programmes using mid-circuit measurement in Guppy; investigations of active-space selection and quantum-selected configuration interaction (QSCI) for the chromium dimer; and the use of the atomic valence active space (AVAS) approach to identify active spaces and perform resource estimation for myoglobin.
One participant reimplemented their research on efficient state-preparation circuits based on orbital entanglement in InQuanto and compared the performance of those circuits with the platform’s efficient ansatz methods on the Helios emulator. It was rewarding to see participants apply these tools to their own research challenges and explore new approaches to quantum chemistry.
With Support from the UK’s National Quantum Computing Centre
This rewarding week was made possible through the support of the UK’s National Quantum Computing Centre, whose partnership helped bring the programme to life, and by the contributions of colleagues across Quantinuum. The teaching team brought together experts from our quantum chemistry, compiler, and error-correction teams, who developed the course materials, delivered lectures, and led the workshops.
Above all, the programme demonstrated the value of bringing researchers together around a shared foundation, practical tools, and opportunities to learn from one another. We look forward to seeing how participants build on these ideas in their future research.
Interested in future training opportunities, workshops, and community events from Quantinuum? Join QNET to stay connected with the latest updates, resources, and opportunities to engage with the quantum computing community.